EoCoE • Science Challenge
Materials for Energy
Description
Scientific Leader: Edoardo Di Napoli, Alessandro Pecchia
This project aims to upgrade libNEGF into the exascale domain, creating the first freely available library tool able to tackle large-scale calculations of non-equilibrium charge carrier transport. This will be the first tool to fully encompass all the relevant interaction mechanisms in order to obtain realistic simulations of nanoscale optoelectronic devices.
The applications of libNEGF will target 2D heterostructures, traditional III- V multi quantum-wells, nano-electronic device simulations (both 2D and 3D) as well as nano-thermoelectric devices.
The goal is to expand the libNEGF library to create a versatile quantum simulation tool that can handle industry-relevant simulations of optoelectronic hetero-structures based on 2D materials with DFT level accuracy. This includes accounting for light-carrier interactions, electron-phonon scattering, and excitonic effects. To achieve this, the libNEGF code must be made scalable on pre-exascale and future exascale systems by developing new algorithms, data distribution methods, and platform-aware kernels.
The NEGF formalism is a computationally intensive method dealing in essence with large dense tensors in configuration, energy, and momentum space. To address the memory scaling issue, strategies such as data distribution, data pipelining, overlapping of communication and computation, and reduced-precision storage of intermediate results must be implemented.
We aim introducing further layers of parallelization and distribution, develop a flexible block-dense structure scheme, consider the effect of data compression and accuracy, and design a new data flow strategy that maximizes the interleaving of computation with communication. In addition, we aim at achieving multi-level parallelism and interoperability between different platforms, such as NVIDIA, Intel, AMD, ARM, etc., by developing an automatic framework that combines compiler-based code generation with heuristic performance prediction tools that evaluate the generated code through modeling of the computation and communication patterns of the underlined algorithm.
LibNEGF is already interfaced to DFT codes such as dftb+ or the finite-element device simulator tiberCAD. Additional interfaces are in progress, with BigDFT with a wavelet or LCAO basis sets, or tools exploiting maximally-localized Wannier representation of the local basis.
To validate the tool’s capabilities and ability to perform at exascale, it will be tested on a few identified structures, such MoS2/WS2 multilayers and MoS2/graphene with varying compositions and transport orientations. The tests will focus on exciton splitting ability and internal quantum efficiencies, representing the first demonstration of NEGF simulations on large-scale hetero-structures of this type.
From HPC Research to Energy Applications
Results
The most important aspect of the Material for Energy scientific challenge of EoCoE is to join scientific and technical competences from the most important institutions and research centres in Europe in order to discover novel materials that allow the production of electricity by harvesting energy from sunlight, salinity gradients and temperature gradients. In order to achieve this challenging purpose, we are developing codes based on theories at different length scales able to be run efficiently on the next generation of supercomputers that are reaching exascale computing performance, i.e., supercomputers able to perform one billion billion (1018) floating-point operations per second.
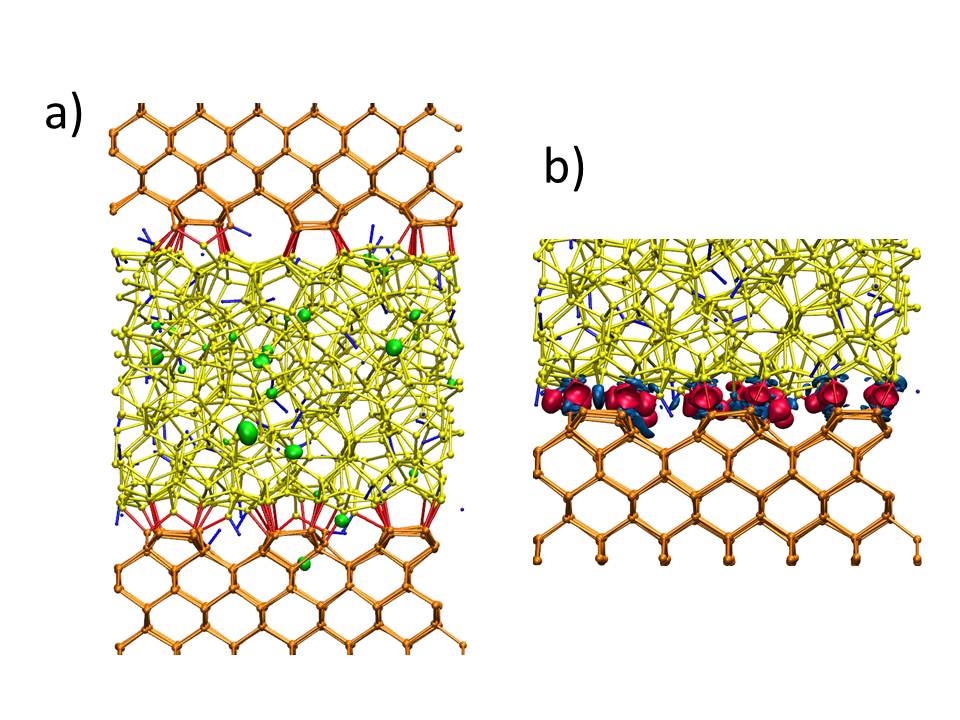
Thanks to the EoCoE project, four different European Institutions (CIEMAT, CNR, ENEA and FZJ) collaborated in understanding the transport mechanisms underlying photovoltaic devices based on the Silicon Heterojunction technology. Amorphous-crystalline heterointerfaces play a crucial role in the photovoltaic operation of the devices, but the microscopic mechanisms of transport and recombination mechanisms at the interface are not yet well understood. The main purpose here is to design and develop cutting-edge computational methods and HPC-ready software for photovoltaic device simulations at atomistic resolution in the most insightful and predictive way. The results look very promising for further research since we have simulated the transport at the interface between passivated Silicon and amorphous Silicon by taking into account the presence of defects in a realistic structure, and increased the speed of specific algorithms used by the code more than six times. Moreover, EoCoE has signed a MoU with MAX CoE to collaborate on the optimization of the pre-processing of the electronic properties (using QUANTUM ESPRESSO) before inserting the definition of these properties in the transport calculation.
Another objective reached by the collaboration of CEA and Maison de la Simulation is to assess the best electrode/electrolyte combination to optimize the electricity production in electrochemical systems. Electrical power production from salinity gradients harvests the free energy lost during the mixing of sweet water with salty water in river estuaries. The main technologies developed for this purpose to date exploit the electric potential differences applied across membranes. We analyse an alternative approach based on capacitive mixing. As part of this approach, we are investigating the electrode structure which optimizes a blue energy production, and which is the compositions that will enable optimal performances of thermo-electrochemical cells. Five-hundred structures never seen before were passed through the artificial intelligence model to predict the vertical energy gaps between the reduced and the oxidized species, necessary to predict electrochemical properties (e.g. redox potentials). Such a tool has the potential to accelerate the research of the best electrode/electrolyte combination to optimize the energy harvesting process.
Working with the University of Bath (research group of Prof. Alison Walker), we have been able to achieve good results on optoelectronic devices based on two classes of emerging photovoltaic materials, namely organic molecular semiconductors and metal-halide perovskite semiconductors. Both types of cells are cheap to make but have some stability concerns. Organic solar cells are only 12% efficient whereas perovskite solar cells are close to 23% efficiency. However, perovskite cells are notorious for poor stability and current-voltage hysteresis. Our purpose is to design and develop HPC ready software for device models of both cell types extending the simulations to systems of several thousands of atoms to understand degradation mechanisms and find materials and architectures that enhance stability.
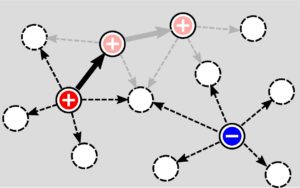
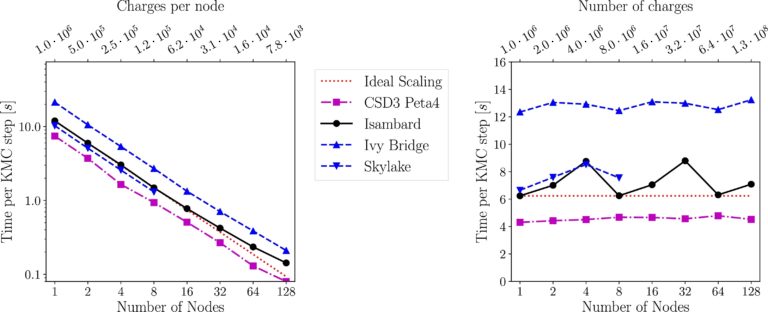
The specificities of these two materials necessitate different computational approaches. Therefore, the development, implementation and application of simulation methodologies amenable to studying phenomena at a broad range of length and time scales has evolved. We are developing a pair of Monte-Carlo codes (KMC and DMC). Notably, algorithmic improvements and other optimization work on the KMC code enabled access to organic molecular semiconductors models that contain millions of charges with good parallel scaling. We demonstrated good performance scaling for 128 million charges up to 128 nodes on Intel and ARM architectures, making possible to model doped organic semi-conductor containing 20,000 charges, while the previous state of the art code, using approximate electrostatics, could perform KMC simulations with systems containing less than 100 charges.
Metal-halide perovskites are currently attracting intense interest for application in optoelectronic devices, due to their remarkable efficiencies in spite of low temperature synthesis methods. In order to test the influence of large polarons on the value of phenomenological parameters, we derived expressions for scattering of polarons by ionised impurities and acoustic phonons, and implemented them in the DMC simulator in order to compute and compare the temperature dependent mobilities of polarons and band electrons in three metal–halide perovskites. We investigated the role played by polaron formation and we found that the main effect is to reduce the mobility of electrons, but the temperature dependence is not altered. This is important for understanding the degradation mechanisms and for the design of solar cells based on metal-halide perovskites with enhanced stability in the performance of the device.
The figures below are snapshots of the results published in scientific articles. Please refer also to the publications section for more details.
Scientific publications related to the Wind for Energy challenge.
Publications
All the EoCoE-II publications are available here
Work in Progress
This section is currently being updated. Please check back soon for more information.